Rev Bras Oftalmol.2026;85:e0087
Choroideremia initially misdiagnosed as retinitis pigmentosa: a multimodal and genetic diagnostic reappraisal

DOI: 10.37039/1982.8551.20260087
ABSTRACT
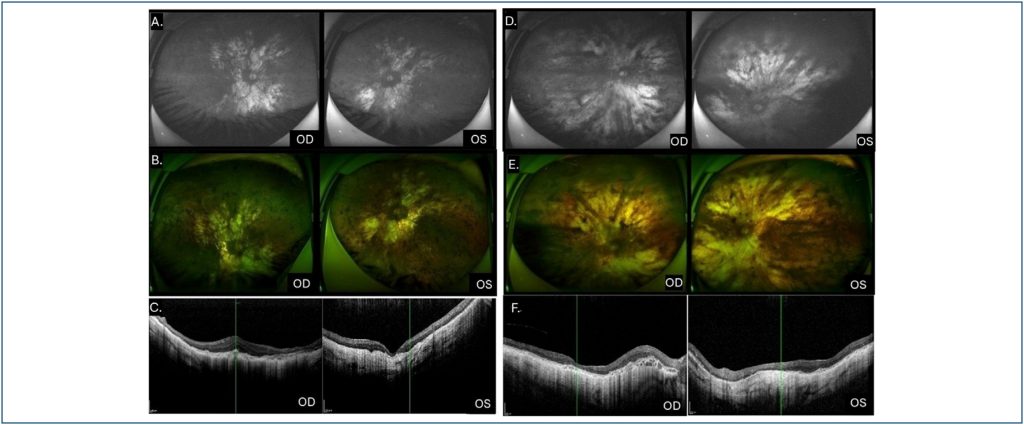
Choroideremia is a rare X-linked chorioretinal dystrophy that may closely resemble retinitis pigmentosa (RP) in its early stages. We describe two brothers who were initially diagnosed with RP and later found to have choroideremia after multimodal imaging and genetic testing. The proband, a 49-year-old man, presented with progressive visual loss, diffuse chorioretinal atrophy, and bone spicule pigmentation. His 51-year-old brother had advanced disease with no light perception. Both displayed fundus autofluorescence and OCT findings typical of choroideremia. A hemizygous nonsense variant in the CHM gene (c.619C>T; p.Arg207*) confirmed the diagnosis. This case highlights the persistent challenge of differentiating early choroideremia from RP and reinforces the importance of genetic analysis in male patients with suspected X-linked retinal dystrophies. Genetic confirmation enables accurate counseling, prognosis, and consideration for emerging gene-based therapies.